cdskit plot - kfuku52/cdskit GitHub Wiki
cdskit plot
cdskit plot renders codon-aware visual summaries from aligned CDS data. The command supports three plotting modes:
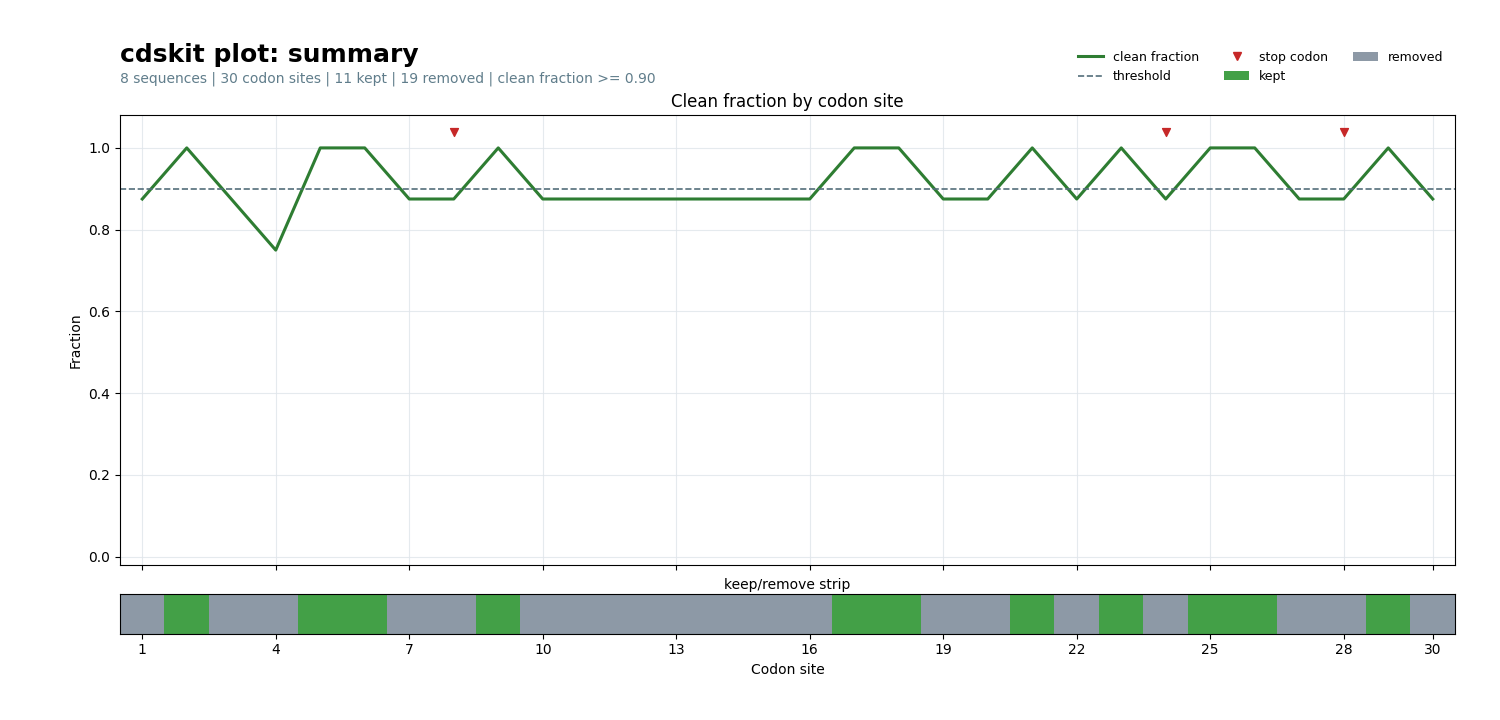
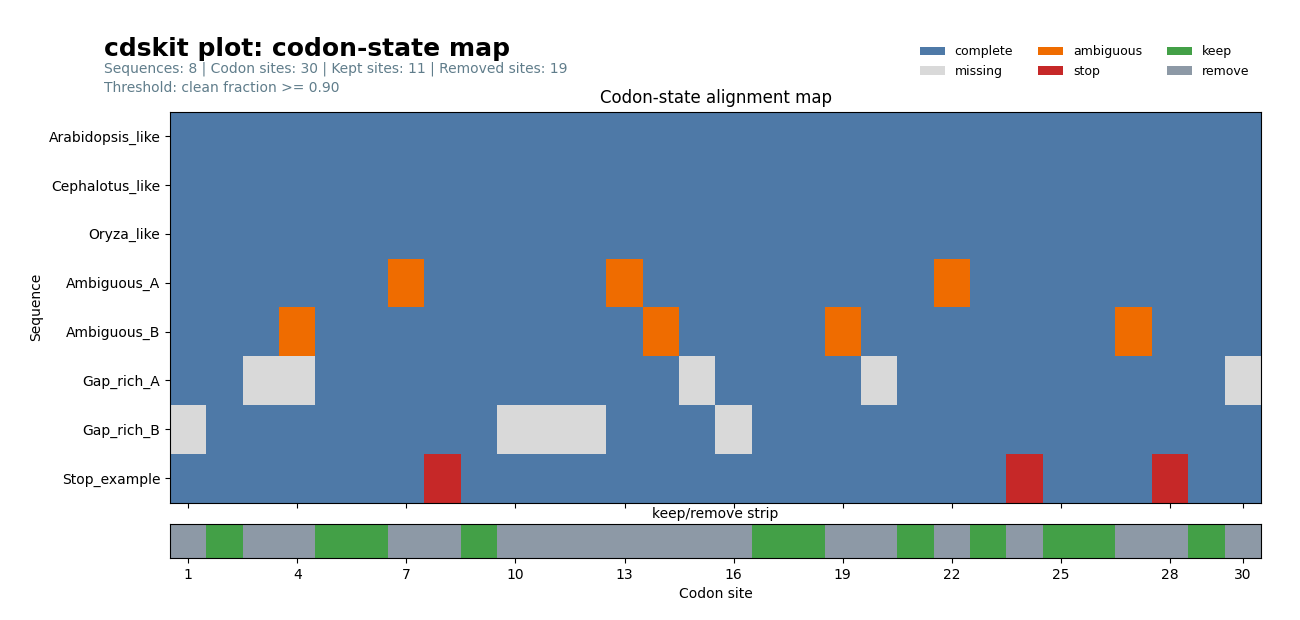
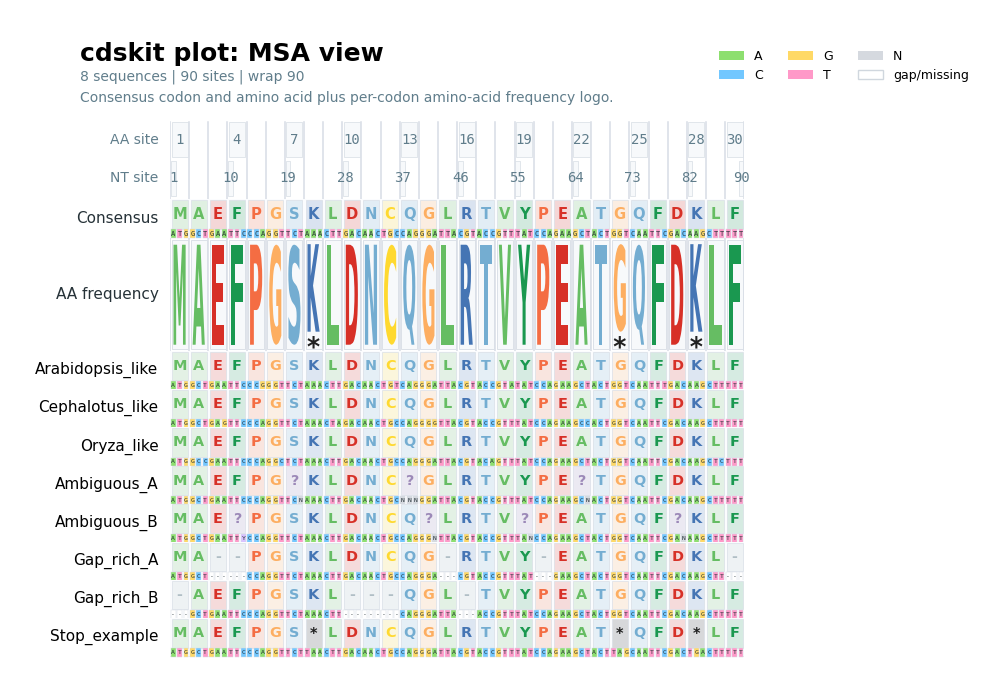
summary: Alignment-level QC summary with a clean-fraction line, threshold overlay, stop-codon markers, and a keep/remove strip.map: Sequence-by-site codon-state map that highlights complete, missing, ambiguous, and stop codons.msa: Nucleotide alignment view with amino-acid site numbers, nucleotide site numbers, consensus amino acid/codon rows, and a per-codon amino-acid frequency logo.
The default output format is PDF. Use --format pdf|svg|png, or let --out_file decide the format automatically from its filename extension.
Examples
The images are saved previews of an illustrative eight-sequence alignment,
with custom figure dimensions. The commands below show the corresponding
modes; your own alignment.fasta determines the plotted values and labels.
Summary plot
cdskit plot --seq_file alignment.fasta --mode summary --out_file qc-summary.pdf --min_clean_fraction 0.9
This mode is useful for choosing a clean-codon fraction threshold before trimming.
Codon-state map
cdskit plot --seq_file alignment.fasta --mode map --out_file codon-map.svg --format svg --min_clean_fraction 0.9
This mode shows how each sequence behaves at each codon site and includes the keep/remove strip derived from the same clean-fraction threshold used by trimcodon.
MSA view
cdskit plot --seq_file alignment.fasta --mode msa --out_file alignment-view.pdf --wrap 90
This mode draws the nucleotide alignment as codon-aware blocks. Each sequence row is split into amino-acid and codon panels, and the consensus plus AA-frequency logo are shown above the alignment. --wrap must be a multiple of three; the preview uses 90, while the default is 60.
Common options
--out_file PATH: Output plot path. Use-for standard output.--format auto|pdf|svg|png: Output format. Ifautois used, the format is inferred from--out_file; otherwise PDF is used by default.--width INT,--height INT: Figure size in pixels.--title STR: Optional plot title.--codon_table INT: NCBI codon table ID.--threads INT: Number of worker threads for data preparation.
Threshold-related options
--min_clean_fraction FLOAT: Minimum fraction of sequences with a clean codon required to keep a codon site insummaryandmap.
A clean codon contains no missing character (-, ?, or .), no ambiguous nucleotide (N, R, Y, etc.), and no definite stop codon under ordinary translation. For example, standard-code
TAR is a definite stop despite being ambiguous. Dual-coding A/C/G/T codons
remain clean despite their context-dependent termination.
Layout options
--row_height INT: Row height formapandmsa.--label_width INT: Width reserved for sequence labels inmapandmsa.--wrap INT: Number of nucleotide columns per block inmsa. Must be divisible by three.--top_n INT: Number of high-ambiguity sequences shown in the side bar chart forsummaryandmap. The default is0, which hides the side chart.
Input requirements
- Input sequences must be DNA.
- All sequences must already be aligned.
- Sequence lengths must be multiples of three.
Notes
summaryandmapare useful for alignment QC and trimming decisions.msais designed for compact codon-aware visualization of moderate-length alignments.- The plotting backend uses
matplotlib, so PDF, SVG, and PNG output are available from the same command.
Codon semantics
See codon meaning and uncertainty for definite versus possible stops, dual-coding tables 27/28/31, overlapping ambiguity/stop counts, and report compatibility. These rules describe sequence QC, not gene function.
References
Cite the method or resource actually used and report the CDSKIT version and options.
- Hunter JD (2007). Matplotlib: A 2D Graphics Environment. Computing in Science & Engineering 9:90–95. Paper — Plotting backend. The MSA logo shows amino-acid frequencies, not information content in bits.